← #nieuws

Boehringer Ingelheim takes its first step into gene therapy

The company have together with the UK Cystic Fibrosis Gene Therapy Consortium (GTC), consisting of Imperial College London and the Universities of Oxford and Edinburgh, Imperial Innovations, and Oxford BioMedica (OXB) announced a global collaboration to develop a first-in-class, long-term therapy for patients with cystic fibrosis (CF).

The new partnership brings together the academic partners’ leading expertise in developing gene therapy for CF and OXB’s leading expertise in manufacturing lentiviral vector-based therapies with Boehringer Ingelheim’s capabilities in drug discovery and the clinical development of novel breakthrough therapeutic agents, reports the collaborators.

A possible universal treatment option

The collaboration will focus on a novel approach using a replication-deficient lentiviral vector in an inhaled formulation, to introduce a healthy copy of the CFTR gene into the cells of the lung. This method has demonstrated high gene transfer efficiency and offers the possibility of repeated administration to maintain the therapeutic effect. Gene therapy is the only therapeutic approach to date that can address all CFTR gene mutations, thus potentially offering a universal treatment option.

Three-way partnership

“This novel three-way partnership brings together an unparalleled combination of clinical, scientific, manufacturing and commercial skills in an effort to develop new treatments and make a major contribution to the lives of patients affected by cystic fibrosis,” stated John Dawson, Chief Executive Officer of Oxford BioMedica. “The GTC has been working determinedly for over 15 years to get to this exciting point of forming a partnership with Boehringer Ingelheim, a global pharmaceutical company with respiratory expertise. Our contribution to this partnership reaffirms our leading position in the development and manufacturing of lentiviral vector gene therapy products at large scale. We look forward to working with our new academic and industry partners.”

“Through this collaboration, we are joining forces with some of the top talents in this disease space to propel treatment advances forward,” said Clive R. Wood, Ph.D., Senior Corporate Vice President, Discovery Research at Boehringer Ingelheim. “Bringing together our existing expertise as a leader for nearly a century in the discovery and development of therapies that have advanced patient care in respiratory diseases with the gene therapy knowledge of our partners, we aim to unlock unprecedented opportunities for patients with this devastating disease, who are desperately waiting for better treatment options.”

Collaboration details

Boehringer Ingelheim has received an option to license the exclusive global rights to develop, manufacture, register, and commercialise this lentiviral vector-based gene therapy for the treatment of cystic fibrosis. Financial terms are not disclosed. During the option period the partners will work closely together to pursue the development of this innovative approach financed by Boehringer Ingelheim. The new collaboration is a joint initiative of Boehringer Ingelheim’s Respiratory Therapeutic Area and Research Beyond Borders (RBB), which is one of the pillars of Boehringer Ingelheim’s research and development strategy and explores emerging science, disease areas and technology. It builds on Boehringer Ingelheim’s commitment to early science and its comprehensive approach to respiratory research and development to investigate new treatments that have significant clinical value in areas of high unmet need.
Source: Nordic life science

← #nieuws

Adaptive pricing: van woorden naar daden

, ,

Mooi nieuws: adaptive pricing van geneesmiddelen, waarbij de hoogte van vergoeding aan de onzekerheid en grootte van het effect aangepast wordt, wint aan draagvlak. In een interview in het NRC noemt niemand minder dan Martin van der Graaff (secretaris WAR, ZIN) adaptive pricing een logische regeling. HollandBIO popelt dan ook om deze oplossing uit haar “Sneller, Beter-programma” samen met het Zorginstituut en andere stakeholders verder uit te werken.

Vergoedingsbeslissingen leiden steeds vaker tot verhitte discussies. Vooral bij weesgeneesmiddelen is het keer op keer raak. Juist door de kleine patiëntengroepen is het voor weesgeneesmiddelen doorgaans lastig om aan de door het Zorginstituut gevraagde bewijslast te voldoen. En zolang er onzekerheid bestaat over de grootte van het effect, is vergoeding en toegang voor nieuwe patiënten geen uitgemaakte zaak. Een vicieuze cirkel, die duidt op een systeem dat vastloopt.

Adaptive pricing heeft alles in zich om deze impasse te doorbreken. Vanwege de kleine populaties en het minimaal beslag dat zij leggen op de zorgkosten (0,24% voor alle weesgeneesmiddelen totaal[1]) ziet HollandBIO in weesgeneesmiddelen de ideale proeftuin. Laten we daarom de handen ineen slaan en werk maken van deze oplossing waarbij gezondheidswinst, betaalbaarheid en innovatie hand in hand gaat.

[1] Zorginstituut, ´Monitor Weesgeneesmiddelen 2017’, 21 december 2017

← #nieuws

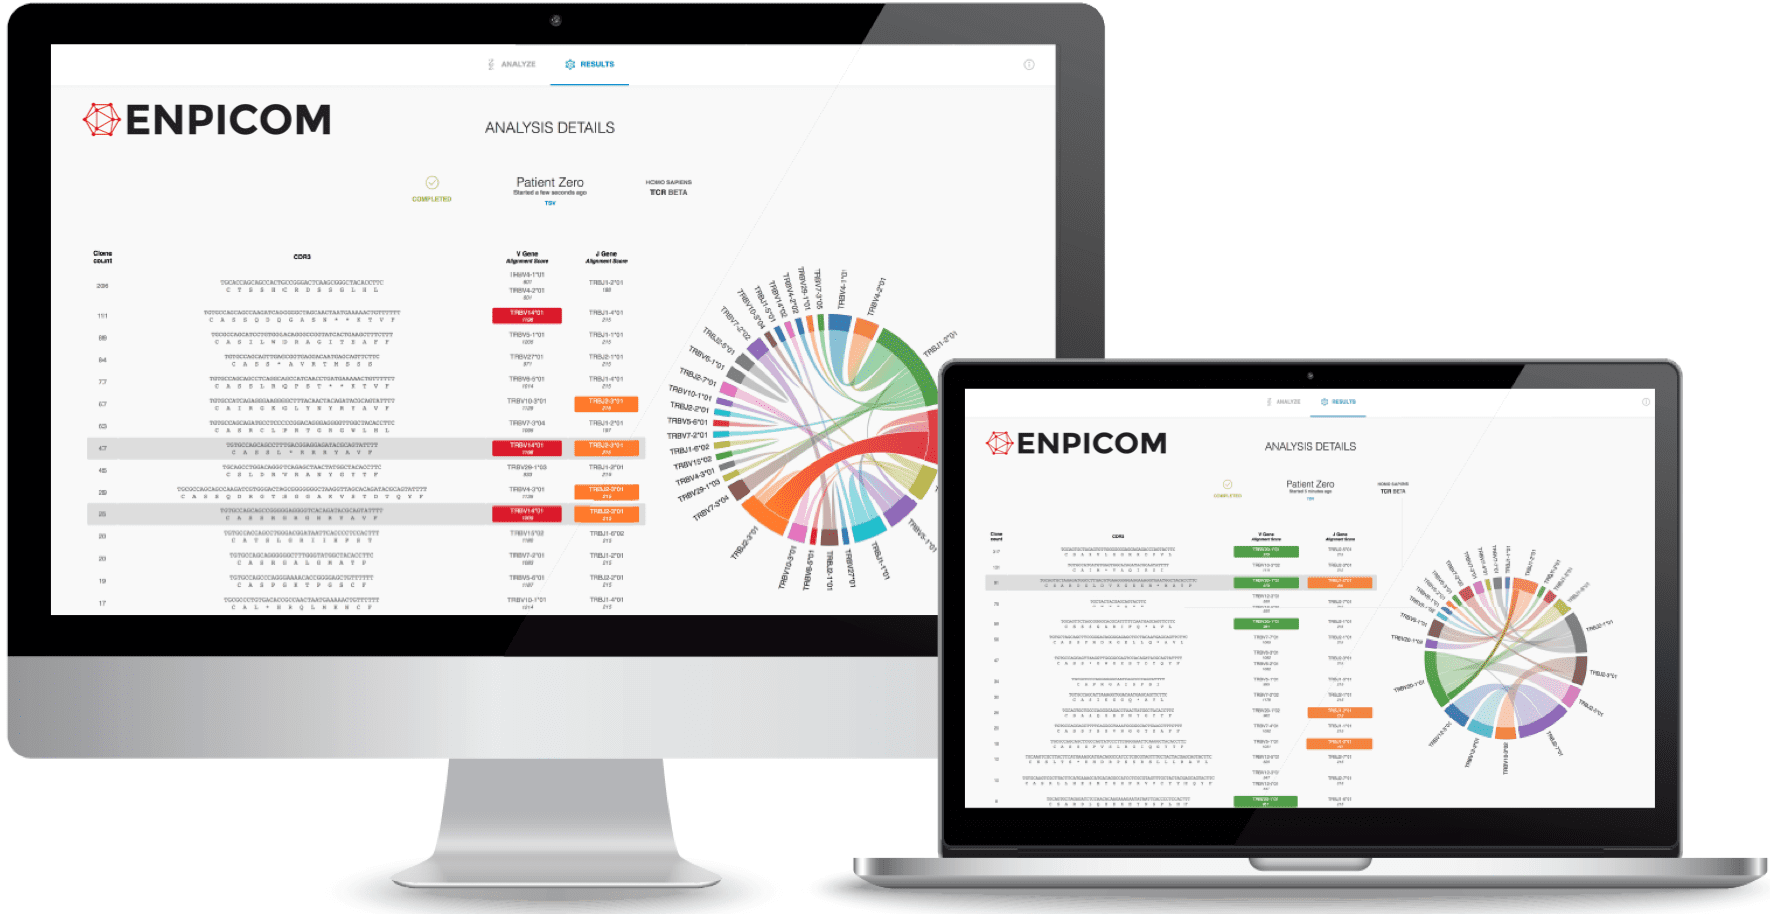
ENPICOM Releases Immune Repertoire Sequencing Analysis Platform to Personalize Immunotherapies

ENPICOM finalized development and released the first version of their ImmunoGenomiX (IGX) platform to support the development, patient stratification and treatment monitoring of immunotherapies.

The adaptive immune system, also known as the acquired immune system, is a subsystem of the overall immune system that is composed of T and B lymphocytes, which eliminate pathogens or prevent their growth. Moreover, the clinical relevance of T cells in the control of different human cancers and autoimmune diseases is beyond doubt. Immunotherapies that boost the ability of T cells to destroy cancer cells have proven therapeutic efficacy in a variety of human malignancies.

ENPICOM develops a T cell and B cell receptor (TCR/BCR) repertoire immunosequencing data analysis solution. This IGX platform will be modularly expanded to become a comprehensive end-to-end immunosequencing data analysis platform designed to analyze, monitor, and compare the immune repertoires in the context of immunotherapy development and at all stages of treatment and disease over time. Starting from high-throughput sequencing data, it will deliver an easy-to-read report depending on the specific application, be it research, diagnosis, patient stratification, or treatment monitoring. The IGX platform will allow customers to use their own sample preparation protocols and the next-generation sequencing technology of choice. It requires no programming skills as the interface is intuitive and flexible.

The first version is available as of today and consists of the base module, IGX Explore, to analyze TCR repertoires from raw sequencing data, report back the number of individual TCR clones and interactively visualize them in a user-friendly way. With IGX Explore (TCR) immunotherapy companies can boost their target discovery, research and drug development processes. In case they need TCR repertoire analysis for a specific application or results visualized in a different way, ENPICOM can custom-build it.

Using a recently published and independent benchmark, ENPICOM showed that IGX Explore offers superior accuracy. The independent benchmark was published in a recent paper by Afzal et al., in which the authors benchmarked ten state-of-the-art TCR repertoire analysis tools. More information about this benchmark study can be found at the company’s website.

Dr. Nicola Bonzanni, Chief Scientific Officer, explains: “An important step in repertoire analysis is the correction of sequencing errors. If sequencing errors were left uncorrected, the richness of a sample would be vastly overestimated; correcting too many errors would lead to under-estimating richness. A critical challenge in immune repertoire analysis is to correct the sequencing errors in the data without losing repertoire-specific information in the process. IGX Explore uses an Adaptive Error Correction algorithm that learns sample-specific mutation rates from the data. By using sample-specific error estimations, erroneous sequences can then be corrected to provide accurate clone sequences, sizes, and frequencies” and adds “These results are very comforting, as highly accurate clonality analysis is the stepping stone towards successful application of this technology in the clinic at later stage”.

← #nieuws

Khondrion receives €2.3M funding from EU Horizon 2020 SME instrument

Khondrion was awarded €2.3M from the highly competitive EU Horizon 2020 SME instrument program. The grant will be used for the execution of a phase 2b dose-finding clinical trial with KH176 in adults with mitochondrial disease and the delivery of the milestones allowing for financing of the pivotal Phase 3 clinical trial for market approval.

Khon2btreat

Khondrion’s project KHON2bTreat “Late Clinical Development of KH176: an innovative orphan drug to reach mitochondrial disease patients & market” is one of the 65 granted projects selected out 1664 proposals.

Jan Smeitink, Khondrion’s CEO, said “This H2020 SME instrument grant is a great opportunity for Khondrion enabling us to accelerate the clinical development of KH176 for patients suffering from mitochondrial disease who have an unmet medical need for clinical relevant treatments. We are extremely proud to be awarded with the most competitive grant for SME’s in Europe and are eager to start and bring this innovative project to a success.”

KH176

KH176 is an orally bio-available small molecule in development by Khondrion for the treatment of mitochondrial (-related) diseases. The compound is a member of a new class of potential Khondrion drugs essential for the control of oxidative and redox alterations. Khondrion reported earlier that KH176 was well tolerated, appeared safe and provided first evidence of efficacy in the KHENERGY study, a Phase II exploratory trial with oral KH176 in the m.3243A>G multisystem mitochondrial MELAS and MIDD syndromes and mixed phenotypes.

← #nieuws

SelectMDx-studie toont kosteneffectiviteit aan in Amerikaanse Gezondheidszorg

MDxHealth meldt de publicatie in The Journal of Urology van een studie die de kosteneffectiviteit van SelectMDx® voor prostaatkanker valideert. SelectMDx® is een niet-invasieve ‘vloeibare biopsie’ test die helpt bij het identificeren van patiënten met een verhoogd risico op agressieve prostaatkanker.


De studie toonde aan dat het opnemen van SelectMDx in het Amerikaanse gezondheidszorgsysteem kan resulteren in een gemiddelde volledige gezondheidswinst van 0,045 levensjaren, met een kostenbesparing van $1,694 per patiënt. Extrapolatie van deze gegevens naar een voorzichtige schatting van 311.879 patiënten per jaar die een biopsie ondergingen, resulteerde in 14.035 levensjaren in volledige gezondheidswinst met een jaarlijkse kostenbesparing van meer dan $500 miljoen voor ieder cohort.


De studie, ontworpen voor het verkrijgen van inzicht in de kosteneffectiviteit van SelectMDx in een populatie van Amerikaanse mannen met een verhoogd PSA-gehalte, laat een model zien met de impact van het gebruik van de niet-invasieve diagnostische test om mogelijk onnodige prostaatbiopsie uit te sparen. Het primaire doel was om veranderingen in gezondheidsresultaten in kaart te brengen, gemeten in voor kwaliteit gecorrigeerde levensjaren. De secundaire doelstelling was het evalueren van de zorgkosten vanuit het perspectief van de Medicare verzekeraar.


Dr. Matthew Resnick van de afdeling urologische chirurgie en gezondheidsbeleid van het Vanderbilt University Medical Center, Nashville, TN zegt: “Deze studie toont aan dat routinematig gebruik van SelectMDx als leidraad voor biopsiebeslissingen de gezondheidsresultaten verbetert en de kosten voor gezondheidszorg in verband met prostaatkanker-risicobeoordeling in een populatie van Amerikaanse mannen verlaagt. Wij zijn van mening dat deze strategie de waarde van conventionele risicobeoordelingsstrategieën vergroot door verbeteringen in zowel kwaliteit als kosten, de teller en noemer van de gezondheidswaarde-vergelijking.”


“Deze overtuigende studieresultaten zijn een verdere validatie van de positieve impact van SelectMDx voor het verlagen van de kosten voor de gezondheidszorg door ondersteuning in de begeleiding van behandelbeslissingen voorafgaand aan prostaatbiopsie,” zegt Dr. Jan Groen, Chief Executive Officer van MDxHealth. “Deze gezondheids- en economische waarde van vroegtijdige toepassing van SelectMDx tijdens het behandelingsproces, zullen onze onderhandelingen met Amerikaanse ziektekostenverzekeraars belangrijk ondersteunen.”


Dr. Groen vervolgt: “Dit is een belangrijke mijlpaal voor onze SelectMDx-test en versterkt onze strategie om verder voet aan de grond te krijgen in de markt voor eerstelijnszorg. Wij zijn van mening dat met deze uitbreiding de markt voor SelectMDx in de VS en Europa kan verviervoudigen tot meer dan twee miljoen patiënten per jaar”.


← #nieuws

Innovative digital pathology network will transform microscopy

,

Philips and Oxford University Hospitals NHS Foundation Trust (OUH), plan to create a digital pathology network to help drive faster and more efficient diagnoses for patients. OUH will deploy the Philips IntelliSite Pathology Solution at the John Radcliffe Hospital in Oxford, which will serve as a central laboratory service for partner sites at Milton Keynes University Hospital NHS Foundation Trust and Great Western Hospital NHS Foundation Trust in Swindon.

Pathologists play a critical role in disease detection, particularly with cancer diagnosis. Traditionally, pathologists analyze tissue samples on glass slides under a microscope. When seeking a consultation with a sub-specialist, these glass slides must be transported to the second site, which can result in lost or damaged slides or delays in diagnosis. By creating a digital network, OUH will leverage the Philips IntelliSite Pathology Solution, including its Ultra Fast Scanner and Image Management System, to allow clinicians across the three regions and within the Thames Valley Cancer Network to collaborate remotely on patient cases. This enhanced collaboration will help reduce delays in slide transport times, encourage more efficient workflows across the sites, and enable quicker access to specialist pathology opinions. Digital pathology will also support NHS Improvement’s proposed Pathology networks, bringing together clinical expertise with the goal of enhancing patient outcomes.

Digital pathology will transform microscopy

“As an NHS Global Digital Exemplar committed to improving patient care by embracing the latest digital technologies and cross-site collaborations, this partnership aims to modernize patient care and offer innovative world-leading services,” said Professor Clare Verrill, Honorary Consultant in Cellular Pathology, Oxford University Hospitals. “Initially starting with some specialist areas, we hope to soon make our pathology services fully digital, meaning our laboratory teams can maximize efficiency and  focus on analysing samples rather than spending time manually transporting slides between hospitals.”

“With 70% of NHS diagnoses requiring a pathology sample, and with sample analysis becoming increasingly sophisticated while demand increases, Philips is committed to collaborating to modernize UK pathology services,” said Marlon Thompson, General Manager of Philips Digital Pathology Solutions. “Philips wants to help providers meet growing demand by moving pathology from the era of microscopes and fragile stacks of sample slides to one of clinical efficacy with sample images uploaded quickly and analysed robustly within the Philips IntelliSite Pathology Solution. I believe this agreement creates the first digital pathology network with scanning capabilities at multiple sites within the NHS in England, which is a great achievement and a very positive prospect for patients and clinicians alike.”

Philips IntelliSite Pathology Solution is an automated digital pathology image creation, viewing and management system comprised of an Ultra Fast Scanner and Image Management System. This solution contains advanced software tools to manage the scanning, storage, presentation, reviewing, and sharing of images. Philips IntelliSite Pathology Solution is available in various countries globally for primary diagnostic use. In addition to its CE-IVD clearance in Europe, it was the first – and currently only – digital pathology solution allowed to be marketed for primary diagnostic use in the U.S. since April 2017 and in Japan since December 2017.

Clinical cases will commence once network installation has completed, expected in Q3 2018.

Source: Phillips

← #nieuws

Biogen en overheid zijn het eens: SMA patiënten krijgen Spinraza

,

Biogen en de Nederlandse overheid hebben een overeenkomst bereikt over de vergoeding van Spinraza. HollandBIO is daar heel blij mee. Spinraza is het eerste en enige goedgekeurde medicijn voor de behandeling van spinale musculaire atrofie (SMA), een zeer ernstige, progressieve spierziekte. HollandBIO is er trots op een sector te mogen vertegenwoordigen die deze innovaties voortbrengt.

Door de overeenkomst komt het middel Spinraza voor circa 80 jonge kinderen met SMA vanaf 1 augustus in het basispakket. Dit is geweldig nieuws voor patiënten en hun ouders. Dorota Mazurkiewicz, Managing Director van Biogen Nederland licht de overeenkomst toe:

“Allereerst is dit prachtig nieuws voor de betreffende SMA-patiënten en hun families. SMA is een verschrikkelijke ziekte waar geen medicijn voor bestond, maar nu kunnen we patiënten nieuwe hoop bieden. We gaan nu met betrokken partijen in gesprek om de mogelijkheden voor voorwaardelijke vergoeding te verkennen. We hopen daarmee brede toegang voor alle SMA-patiënten te kunnen realiseren, ongeacht leeftijd of ziektetype, zoals ook het geval is in veel andere landen.”

We zijn er dus nog niet. Het middel betekent een kentering in het leven van patiënten. HollandBIO hoopt daarom dat alle patiënten met SMA zo snel mogelijk toegang krijgen tot Spinraza.

← #nieuws

Galapagos announces design for PINTA Phase 2 trial with GLPG1205 in IPF

Galapagos announces the PINTA Phase 2 trial design with its GPR84 inhibitor GLPG1205 in patients with idiopathic pulmonary fibrosis (IPF). PINTA is a randomized, double-blind, placebo-controlled trial investigating a 100 mg once-daily oral dose of GLPG1205. The drug candidate or placebo will be administered for 26 weeks in up to 60 IPF patients. Patients may remain on their local standard of care as background therapy. Primary objective of the trial is to assess the change from baseline in Forced Vital Capacity (FVC in mL) over 26 weeks compared to placebo. Secondary measures include safety, tolerability, pharmacokinetics and pharmacodynamics, time to major events, changes in functional exercise capacity, and quality of life. IPF diagnosis will be confirmed by central reading. Recruitment for PINTA is planned in 10 countries in Europe, North Africa, and the Middle East. First dosing of an IPF patient is expected in the second half of 2018.

GLPG1205 is a GPR84 inhibitor discovered by Galapagos and fully proprietary to Galapagos. GLPG1205 showed a reduction in signs and symptoms in IPF animal models and has shown favorable tolerability in healthy volunteers and ulcerative colitis patients in previous trials. Galapagos currently has three drug candidates with distinct mechanisms of action in its fully proprietary portfolio aimed at building an IPF franchise: GLPG1690 in the ISABELA Phase 3 program, GLPG1205 in PINTA Phase 2, and GLPG3499, currently in pre-clinical development.

“GLPG1205 has shown signs of good activity in relevant animal models, and GPR84 has already been validated as a mechanism in combination with nintedanib[1] in IPF,” added Dr. Piet Wigerinck, Chief Scientific Officer of Galapagos. “We have a well-designed trial with PINTA for ‘1205 that we anticipate will give us new insights into the potential value of GPR84 inhibition as a mechanism to treat this highly fatal disease.”


Source: Galapagos

← #nieuws

First cohort enrolled in trial with innovative gene therapy for arthritis

Arthrogen, a clinical stage gene therapy company together with the Centre for Human Drug Research (CHDR), announced that the first three arthritis patients have been enrolled in a phase Ib gene therapy trial for treatment of arthritis with ART-I02.

ART-I02 is an adeno-associated virus (AAV5) vector encoding the human IFN-β gene under control of an inflammation-responsive promoter. ART-I02 is designed to produce the anti-inflammatory protein IFNβ in the synovial cells in the joint. The aim is to achieve a sustained clinical remission with a single treatment. This current clinical phase Ib trial, evaluates the safety, tolerability, pharmacokinetics, immunogenicity and anti-inflammatory activity of ART-I02 treatment in patients with rheumatoid arthritis (RA) or osteoarthritis (OA). In total 12 patients with RA or OA in their hand joints will be included. Parallel to this study, a similar phase Ib trial has started in Canada treating 15 patients with RA in their wrist.

“Patients with rheumatoid arthritis and osteoarthritis face chronic pain and immobility, even despite the availability of new, mostly systemic treatments.”, explains principal investigator Prof. dr. Koos Burggraaf of CHDR and he continuous: “In this trial we evaluate the safety of ART-I02 and we assess effect measures, hoping that this first step may prove to be an important milestone marking a new era in the treatment of arthritis, a chronic debilitating disease.”.

Robert Jan Lamers, Chief Executive Officer of Arthrogen comments ‘We would like to thank our investors, collaborators and participating patients in helping us achieve this pivotal milestone. Based on anticipated enrollment and follow-up timelines, the first set of data is expected to be available in the fourth quarter of 2018’.

Source: Arthrogen

← #nieuws

Eisai and Biogen announce positive topline results

Eisai and Biogen announced positive topline results from the Phase II study with BAN2401, an anti-amyloid beta protofibril antibody, in 856 patients with early Alzheimer’s disease. The study achieved statistical significance on key predefined endpoints evaluating efficacy at 18 months on slowing progression in Alzheimer’s Disease Composite Score (ADCOMS) and on reduction of amyloid accumulated in the brain as measured using amyloid-PET (positron emission tomography).

Study 201 (ClinicalTrials.gov identifier NCT01767311) is a placebo-controlled, double-blind, parallel-group, randomized study in 856 patients with mild cognitive impairment (MCI) due to Alzheimer’s disease (AD) or mild Alzheimer’s dementia (collectively known as early Alzheimer’s disease) with confirmed amyloid pathology in the brain. Efficacy was evaluated at 18 months by predefined conventional statistics on ADCOMS, which combines items from the Alzheimer’s Disease Assessment Scale-cognitive subscale (ADAS-Cog), the Clinical Dementia Rating Sum of Boxes (CDR-SB) scale and the Mini-Mental State Examination (MMSE) to enable sensitive detection of changes in early AD symptoms. Patients were randomized to five dose regimens, 2.5 mg/kg biweekly, 5 mg/kg monthly, 5 mg/kg biweekly, 10 mg/kg monthly and 10 mg/kg biweekly, or placebo.

Topline results of the final analysis of the study demonstrated a statistically significant slowing of disease progression on the key clinical endpoint (ADCOMS) after 18 months of treatment in patients receiving the highest treatment dose (10 mg/kg biweekly) as compared to placebo. Results of amyloid PET analyses at 18 months, including reduction in amyloid PET standardized uptake value ratio (SUVR) and amyloid PET image visual read of subjects converting from positive to negative for amyloid in the brain, were also statistically significant at this dose. Dose-dependent changes from baseline were observed across the PET results and the clinical endpoints. Further, the highest treatment dose of BAN2401 began to show statistically significant clinical benefit as measured by ADCOMS as early as 6 months including at 12 months.

BAN2401 demonstrated an acceptable tolerability profile through 18 months of study drug administration. The most common treatment emergent adverse events were infusion-related reactions and Amyloid Related Imaging Abnormalities (ARIA). Infusion related reactions were mostly mild to moderate in severity. Incidence of ARIA-E (edema) was not more than 10% in any of the treatment arms, and less than 15% in patients with APOE4 at the highest dose per the study protocol safety and reporting procedures.

Detailed results of the study will be presented at future academic conferences.

“The 18-month results of the BAN2401 trial are impressive and provide important support for the amyloid hypothesis,” said Jeff Cummings, M.D., founding director, Cleveland Clinic Lou Ruvo Center for Brain Health. “I look forward to seeing the full data set shared with the broader Alzheimer’s community as we advance against this devastating disease.”

“This is the first late-stage anti-amyloid antibody study to successfully achieve statistically significant results at 18 months, further validating the amyloid hypothesis,” said Lynn Kramer, M.D., Chief Clinical Officer and Chief Medical Officer, Neurology Business Group, Eisai. “We will discuss these very encouraging results with regulatory authorities to determine the best path forward. We continue to work towards the goal of delivering BAN2401 to patients and healthcare professionals as early as possible.”

“The prospect of being able to offer meaningful disease-modifying therapies to individuals suffering from this terrible disease is both exciting and humbling,” said Alfred Sandrock, M.D., Ph.D., executive vice president and chief medical officer at Biogen. “These BAN2401 18-month data offer important insights in the investigation of potential treatment options for patients with Alzheimer’s disease and underscores that neurodegenerative diseases may not be as intractable as they once seemed.”

As reported in December 2017, the study did not achieve its primary outcome measure which was designed to enable a potentially more rapid entry into Phase III development based on Bayesian analysis at 12 months of treatment. Upon the final analysis at 18 months using predefined conventional statistical method, the study did demonstrate a statistically significant slowing of disease progression on the key clinical endpoint (ADCOMS) after 12 months of treatment in patients receiving the highest treatment dose (10 mg/kg biweekly) as compared to placebo.

This release discusses investigational uses of an agent in development and is not intended to convey conclusions about efficacy or safety. There is no guarantee that any investigational uses of such product will successfully complete clinical development or gain health authority approval.


Source: Biogen